Publications
Chronological publications in which Ze Zhang was involved. [ZZ* is (co-)corresponding author. ZZ† is (co-)first author.]
Publications by Year
2026
-
 Gelatin pretreatment enables cryosectioning of brittle tissues for spatial transcriptomicsZe Zhang†* , Chunlu Feng† , Liyuan Wang† , Xinlei Cai , Shiyu Mo , Huizhong Chang , Zhenkun Zhuang , Duoyuan Chen , Xinyue Hu , Wenjie Guo , Yi Lu , Song Feng , Xun Xu , Shiping Liu , Minxiao Wang* , and Youning Lin*iScience, 2026Accepted in principle on November 4, 2026
Spatially resolved transcriptomics (SRT) requires high-quality tissue sections that preserve morphology and RNA integrity, yet brittle, mucus-rich, or highly hydrated specimens often fail under standard OCT or FFPE workflows, leading to tissue damage and RNA loss. We present GSP-Cryo (Gelatin Solution Pretreatment Cryosectioning), a practical strategy that enhances tissue mechanical stability through gelatin perfusion or immersion prior to freezing, while remaining compatible with downstream molecular assays. Across specimens with diverse structural properties, GSP-Cryo consistently enabled continuous, morphologically intact cryosections. Using the moon jellyfish Aurelia coerulea, we optimized immersion and permeabilization parameters and validated performance across developmental stages. GSP-Cryo–prepared adult sections supported full Stereo-seq library construction and sequencing, yielding a spatial transcriptomic atlas of the umbrella margin with spatial patterns. By overcoming long-standing barriers to cryosectioning fragile tissues, GSP-Cryo provides an efficient workflow that facilitates spatial transcriptomics in challenging non-model organisms and delicate tissue types.
@article{20261104, author = {Zhang†*, Ze and Feng†, Chunlu and Wang†, Liyuan and Cai, Xinlei and Mo, Shiyu and Chang, Huizhong and Zhuang, Zhenkun and Chen, Duoyuan and Hu, Xinyue and Guo, Wenjie and Lu, Yi and Feng, Song and Xu, Xun and Liu, Shiping and Wang*, Minxiao and Lin*, Youning}, title = {Gelatin pretreatment enables cryosectioning of brittle tissues for spatial transcriptomics}, journal = {iScience}, year = {2026}, month = {}, day = {}, pages = {}, volume = {}, number = {}, doi = {}, url = {}, eprint = {}, note = {Accepted in principle on November 4, 2026}, } -
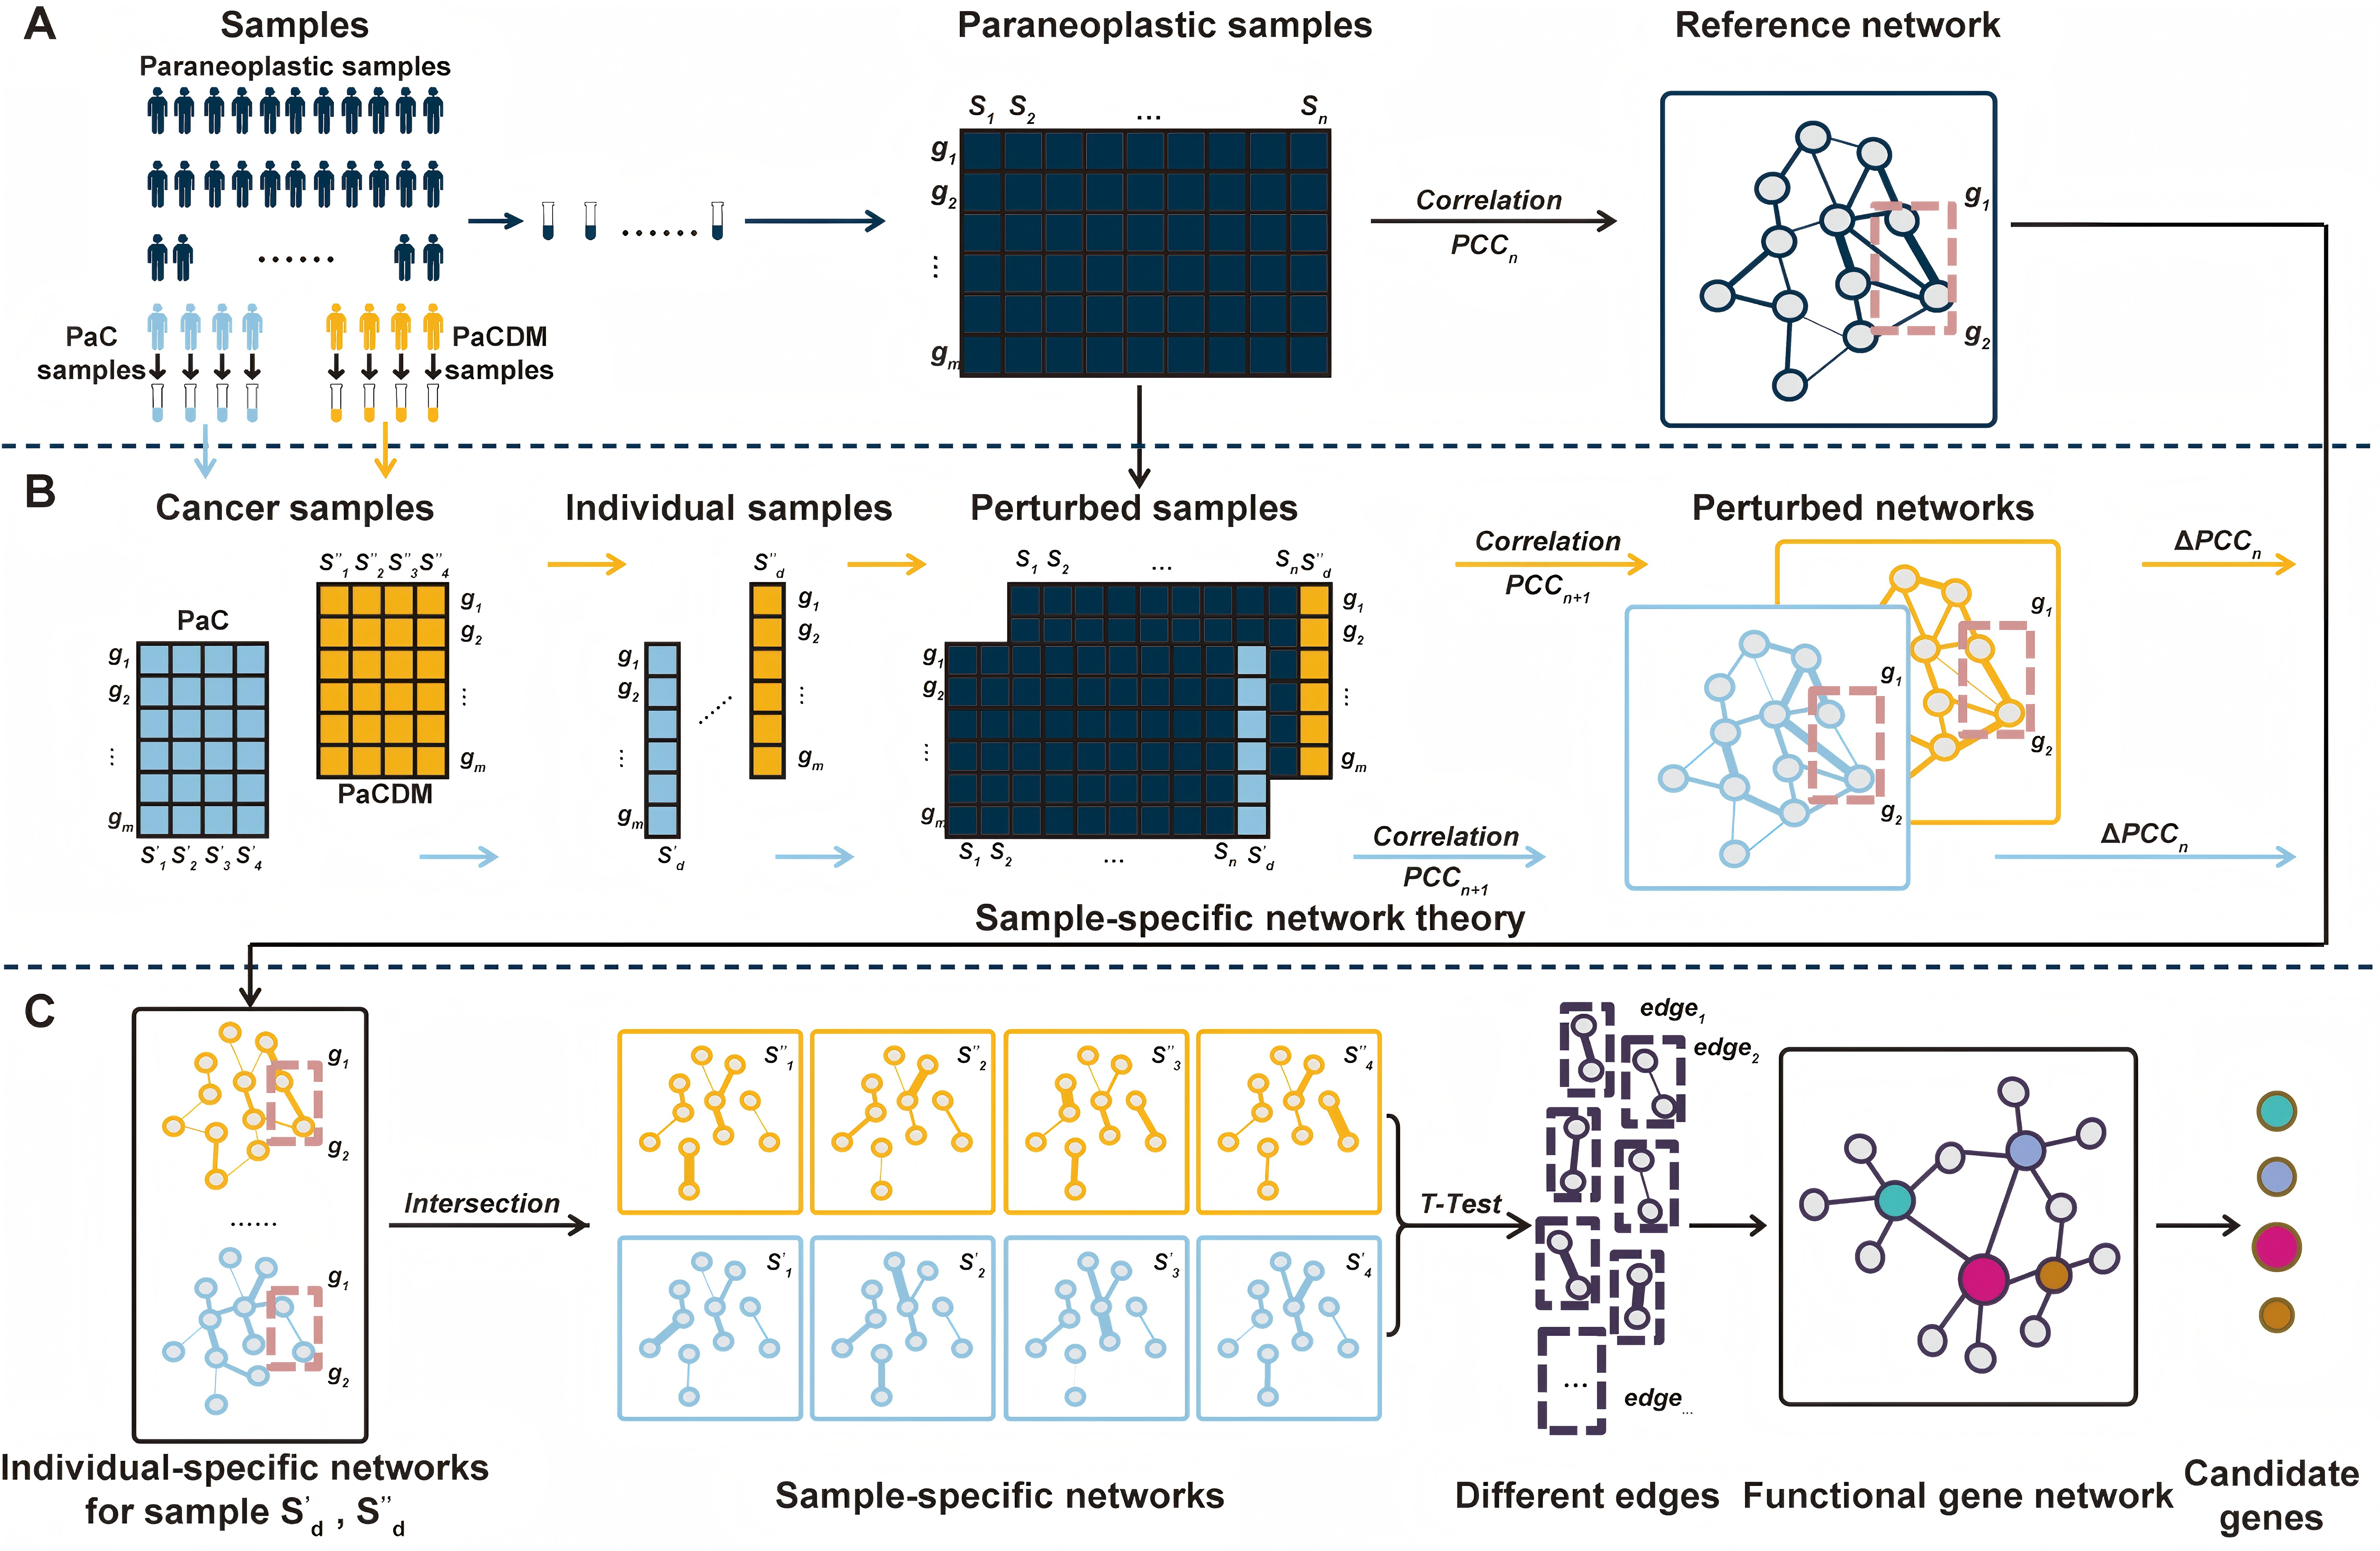 NEK2 and PLK1 in pancreatic cancer cells induce endoplasmic reticulum stress-mediated dysfunction in beta cells under pancreatic cancer-associated diabetes mellitusJianyao Lou† , Ze Zhang† , Zengyu Feng† , Haocheng Yu , Yuheng Tu , Jiali Gong , Yongji Sun , Meihua Shangguan , Kaiyue Pu , Ning Wang , Kaili Cai , Changming Lv , Xiawei Li* , Xiaoping Liu* , and Yulian Wu*TransMed, Nov 2026
Pancreatic cancer-associated diabetes mellitus (PaCDM) mechanisms remain unclear, hindering early detection of pancreatic cancer (PaC) in patients with new-onset diabetes. Using sample-specific network analysis of PaC RNA-seq data and immunohistochemical validation, we identified NEK2 and PLK1 as potential driver genes upregulated in PaC, especially in cases with new-onset diabetes. Overexpression of NEK2 or PLK1 in PaC cells impaired β-cell function, reducing insulin secretion, insulin mRNA/protein, and β-cell viability independently. The NEK2 inhibitor Nek2-IN-5 and PLK1 inhibitor BI6727 reproduced the effects of NEK2 or PLK1 knockdown. Mechanistically, overexpression of NEK2 or PLK1 in PaC cells triggered endoplasmic reticulum (ER) stress, apoptosis, and proinsulin accumulation in β cells; these effects were reversed by the ER stress inhibitor RH01386, and the decreased insulin content was ER stress-dependent. In vivo, silencing NEK2 or PLK1 alleviated glucose intolerance and ER stress in mice with subcutaneous/orthotopic Panc02 tumors. Thus, NEK2 and PLK1 in PaC cells induce ER stress-mediated β-cell dysfunction in PaCDM, and their downstream molecules may serve as targets for early detection of PaC in patients with new-onset diabetes.
@article{10.1016/j.tmed.2026.100068, author = {Lou†, Jianyao and Zhang†, Ze and Feng†, Zengyu and Yu, Haocheng and Tu, Yuheng and Gong, Jiali and Sun, Yongji and Shangguan, Meihua and Pu, Kaiyue and Wang, Ning and Cai, Kaili and Lv, Changming and Li*, Xiawei and Liu*, Xiaoping and Wu*, Yulian}, title = {{NEK2} and {PLK1} in pancreatic cancer cells induce endoplasmic reticulum stress-mediated dysfunction in beta cells under pancreatic cancer-associated diabetes mellitus}, journal = {TransMed}, year = {2026}, month = nov, day = {15}, pages = {100068}, volume = {}, number = {}, doi = {10.1016/j.tmed.2026.100068}, url = {https://doi.org/10.1016/j.tmed.2026.100068}, eprint = {https://doi.org/10.1016/j.tmed.2026.100068}, } -
 SpecBridge: Spectral Structure Alignment and Transitive Bridging for 3D–2D–Text Pre-TrainingDong Wang , Jie Jiang* , Weidong Min , Lixin Zhan , Xinpeng Zhao , and Ze ZhangInternational Joint Conference on Artificial Intelligence 2026 (IJCAI2026), Sep 2026
Open-vocabulary 3D understanding aims to align 3D representations with a unified vision-language semantic space. However, existing methods suffer from the challenge of structural asymmetry caused by sparse observations and holistic geometries. Additionally, the inherent semantic chasm between discrete coordinates and abstract natural language hinders cross-modal mapping. This work introduces SpecBridge, a 3D-2D-Text pre-training framework that leverages CLIP priors as a foundational bridge to connect three modalities by synergizing spectral graph theory with transitive semantic learning. The method consists of two main sub-modules. First, Spectral Eigen-Modal Alignment is presented to construct cross-modal correlations within the intrinsic eigen-spectral space via Laplacian eigendecomposition. By aligning low-frequency geometric harmonics with CLIP-guided observation inference, it maintains structural consistency between 3D geometries and 2D projections. Second, a Transitive Spectral-Semantic Alignment method is developed to establish a 3D→2D→Text propagation chain across the CLIP priors bridge. It distills dense CLIP priors into 3D representations through transitive distillation, effectively mitigating the semantic chasm between geometry and language. Extensive experiments confirm that the proposed SpecBridge demonstrates state-of-the-art performance by overcoming challenges triggered by modality discrepancies.
@article{10.24963/ijcai.2026/192, author = {Wang, Dong and Jiang*, Jie and Min, Weidong and Zhan, Lixin and Zhao, Xinpeng and Zhang, Ze}, title = {SpecBridge: Spectral Structure Alignment and Transitive Bridging for 3D–2D–Text Pre-Training}, journal = {International Joint Conference on Artificial Intelligence 2026 (IJCAI2026)}, year = {2026}, month = sep, day = {16}, pages = {1722--1730}, volume = {3}, number = {ijcai2026p192}, doi = {10.24963/ijcai.2026/192}, url = {https://www.ijcai.org/proceedings/2026/192}, eprint = {https://www.ijcai.org/proceedings/2026/192}, } -
 Chromosome-level genome assembly and annotation of the moon jellyfish Aurelia coeruleaXinyue Hu† , Yang Guo† , Ze Zhang†* , Liyuan Wang , Duoyuan Chen , Zhenkun Zhuang , Minxiao Wang , Shiping Liu , and Song Feng*Scientific Data, Apr 2026
Aurelia coerulea (moon jellyfish), a radially symmetrical metazoan in the phylum Cnidaria, possesses key features pertinent to understanding the evolutionary origins of nervous systems. Here, we employed a combination of long-read sequencing, short-read sequencing, and Hi-C chromatin conformation capture techniques to generate a chromosome-level assembly of the A. coerulea genome. The final assembly comprises approximately 554.10 Mb distributed across 21 chromosomes, achieving a scaffold N50 of 24.06 Mb and demonstrating high completeness (protein BUSCO score: 93.0%). Approximately 71.48% of the genome consists of transposable elements. We identified 26,777 protein-coding genes, of which approximately 72.28% have been functionally annotated. This chromosome-level genome provides an essential resource for elucidating early neural evolution and advancing our understanding of cnidarian biology.
@article{10.1038/s41597-026-07102-9,
  author = {Hu†, Xinyue and Guo†, Yang and Zhang†*, Ze and Wang, Liyuan and Chen, Duoyuan and Zhuang, Zhenkun and Wang, Minxiao and Liu, Shiping and Feng*, Song},
  title = {Chromosome-level genome assembly and annotation of the moon jellyfish Aurelia coerulea},
  journal = {Scientific Data},
  year = {2026},
  month = apr,
  day = {10},
  pages = {854},
  volume = {13},
  number = {1},
  doi = {10.1038/s41597-026-07102-9},
  url = {https://www.nature.com/articles/s41597-026-07102-9},
  eprint = {https://www.nature.com/articles/s41597-026-07102-9},
}


2025
-
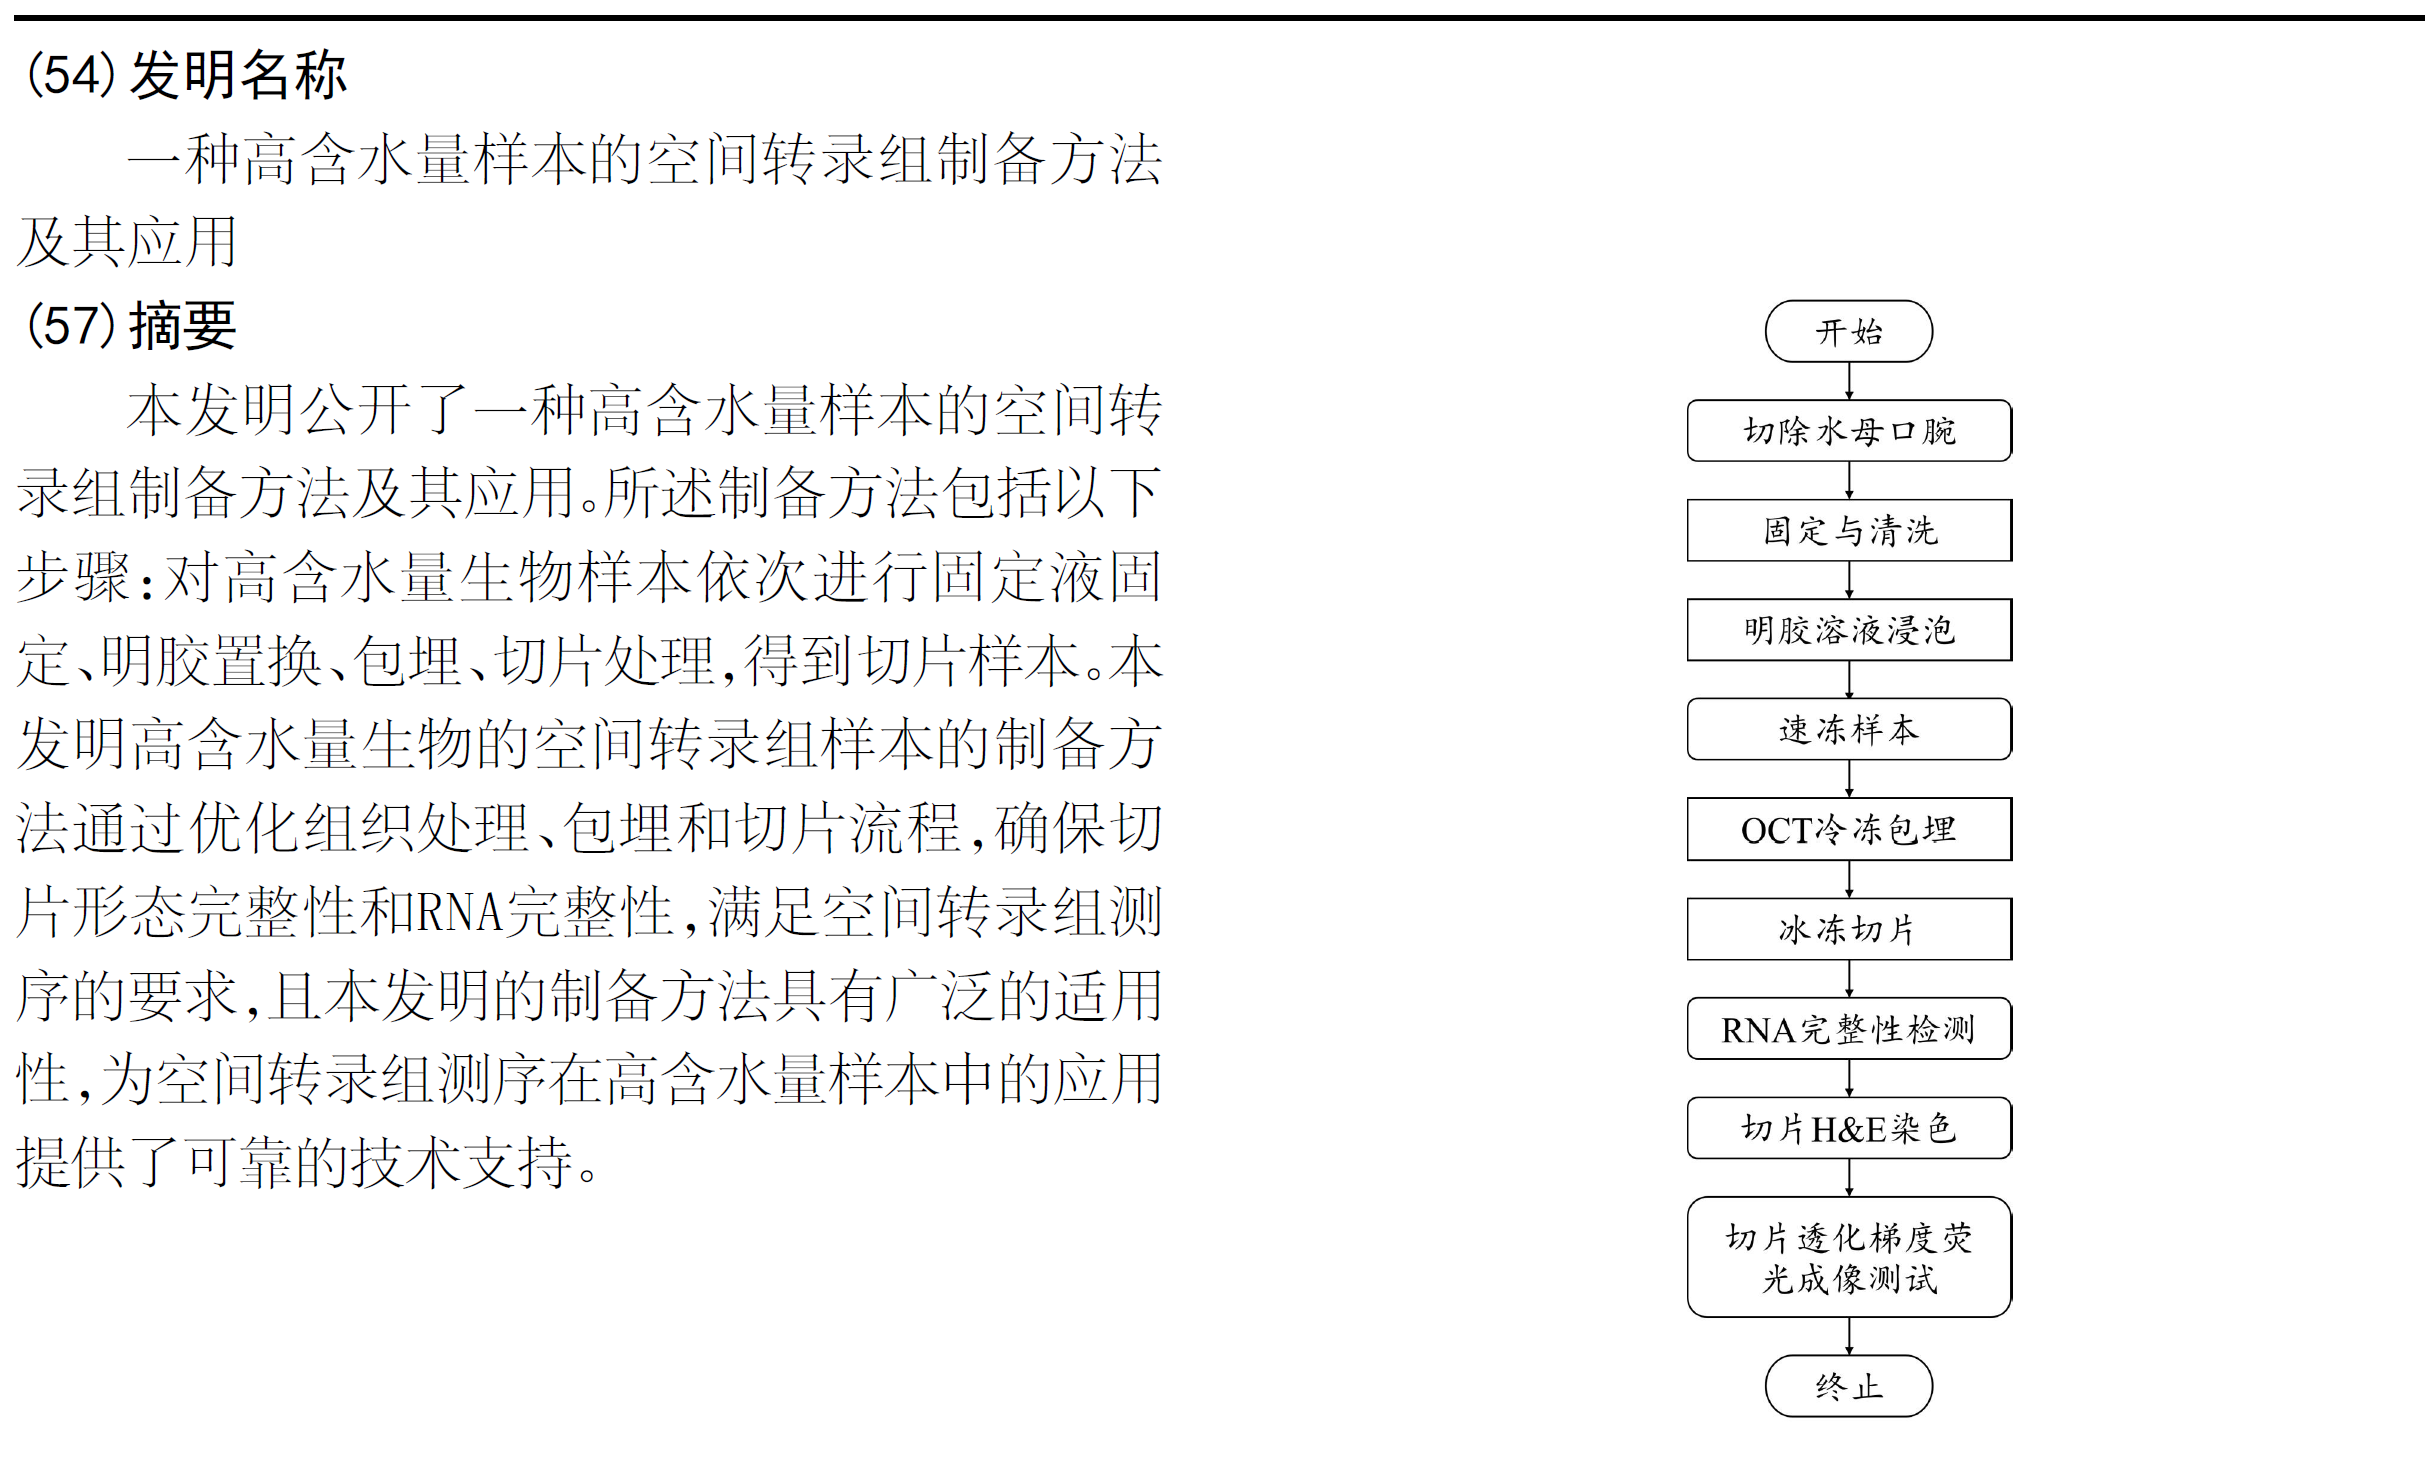 一种高含水量样本的空间转录组制备方法及其应用张泽 , 冯春露 , 庄镇堃 , 陈成 , 胡馨月 , and 陈铎元CN Patent, Nov 2025
本发明公开了一种高含水量样本的空间转录组制备方法及其应用。所述制备方法包括以下步骤:对高含水量生物样本依次进行固定液固定、明胶置换、包埋、切片处理,得到切片样本。本发明高含水量生物的空间转录组样本的制备方法通过优化组织处理、包埋和切片流程,确保切片形态完整性和RNA完整性,满足空间转录组测序的要求,且本发明的制备方法具有广泛的适用性,为空间转录组测序在高含水量样本中的应用提供了可靠的技术支持。
@article{CN120275136B, author = {张泽 and 冯春露 and 庄镇堃 and 陈成 and 胡馨月 and 陈铎元}, title = {一种高含水量样本的空间转录组制备方法及其应用}, journal = {CN Patent}, year = {2025}, month = nov, day = {04}, pages = {}, volume = {G01N1/28; G01N1/30; G01N1/36; G01N1/42; G01N1/34; C12Q1/6806; C12Q1/6869; G01N21/64}, number = {}, doi = {}, url = {https://www.patentguru.com/cn/search?q=CN120275136B}, eprint = {https://www.patentguru.com/cn/search?q=CN120275136B}, } -
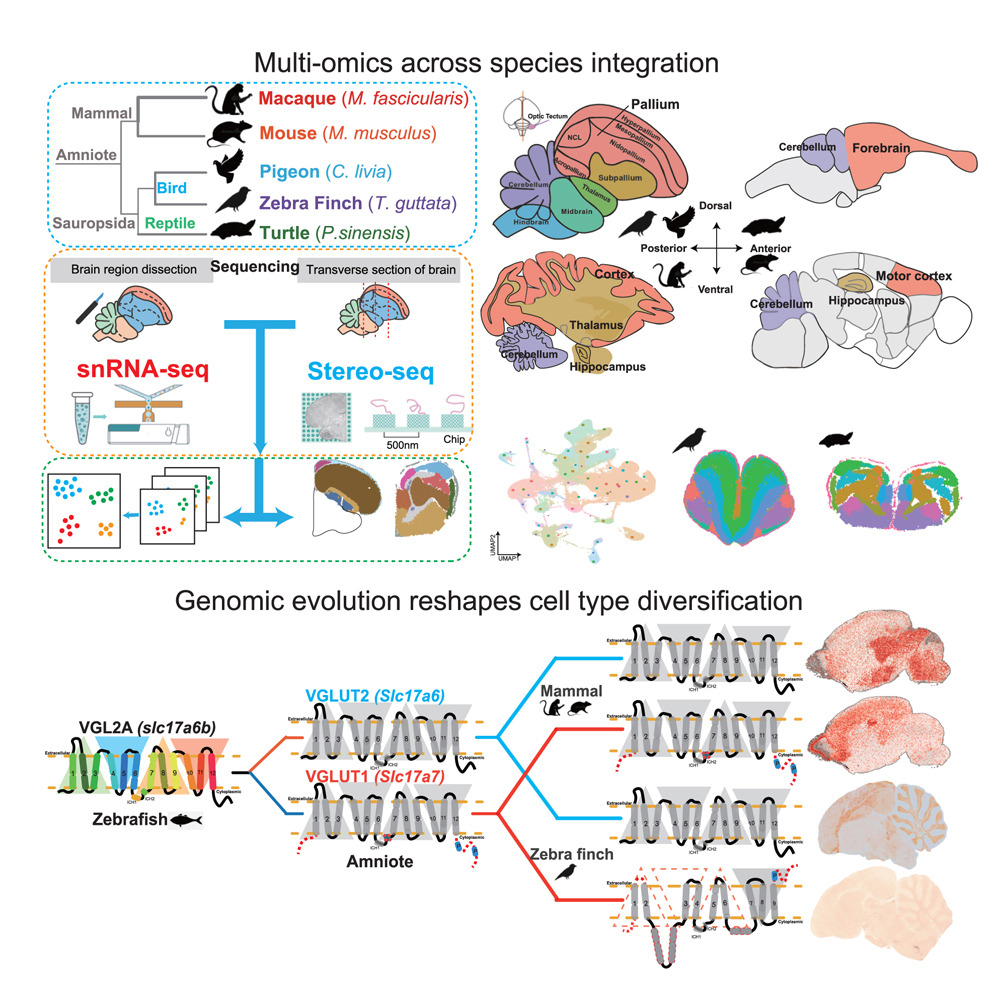 Genomic evolution reshapes cell-type diversification in the amniote brainDuoyuan Chen† , Zhenkun Zhuang† , Maolin Huang† , Yunqi Huang† , Yuting Yan , Yanru Zhang , Youning Lin , Xiaoying Jin , Yuanmei Wang , Jinfeng Huang , Wenbo Xu , Jingfang Pan , Hong Wang , Fubaoqian Huang , Kuo Liao , Mengnan Cheng , Zhiyong Zhu , Yinqi Bai , Zhiwei Niu , Ze Zhang , Ya Xiang , Xiaofeng Wei , Tao Yang , Tao Zeng , Yuliang Dong , Ying Lei , Yangang Sun , Jian Wang , Huanming Yang , Yidi Sun , Gang Cao , Muming Poo , Longqi Liu , Robert K. Naumann* , Chun Xu* , Zhenlong Wang* , Xun Xu* , and Shiping Liu*Developmental Cell, May 2025
Over 320 million years of evolution, amniotes have developed complex brains and cognition through largely unexplored genetic and gene expression mechanisms. We created a comprehensive single-cell atlas of over 1.3 million cells from the telencephalon and cerebellum of turtles, zebra finches, pigeons, mice, and macaques, employing single-cell resolution spatial transcriptomics to validate gene expression patterns across species. Our study identifies significant species-specific variations in cell types, highlighting their conservation and diversification in evolution. We found pronounced differences in telencephalon excitatory neurons (EXs) and cerebellar cell types between birds and mammals. Birds predominantly express SLC17A6 in EX, whereas mammals express SLC17A7 in the neocortex and SLC17A6 elsewhere, possibly due to loss of function of SLC17A7 in birds. Additionally, we identified a bird-specific Purkinje cell subtype (SVIL+), implicating the lysine-specific demethylase 11 (LSD1)/KDM1A pathway in learning and circadian rhythms and containing numerous positively selected genes, which suggests an evolutionary optimization of cerebellar functions for ecological and behavioral adaptation. Our findings elucidate the complex interplay between genetic evolution and environmental adaptation, underscoring the role of genetic diversification in the development of specialized cell types across amniotes.
@article{10.1016/j.devcel.2025.04.014, author = {Chen†, Duoyuan and Zhuang†, Zhenkun and Huang†, Maolin and Huang†, Yunqi and Yan, Yuting and Zhang, Yanru and Lin, Youning and Jin, Xiaoying and Wang, Yuanmei and Huang, Jinfeng and Xu, Wenbo and Pan, Jingfang and Wang, Hong and Huang, Fubaoqian and Liao, Kuo and Cheng, Mengnan and Zhu, Zhiyong and Bai, Yinqi and Niu, Zhiwei and Zhang, Ze and Xiang, Ya and Wei, Xiaofeng and Yang, Tao and Zeng, Tao and Dong, Yuliang and Lei, Ying and Sun, Yangang and Wang, Jian and Yang, Huanming and Sun, Yidi and Cao, Gang and Poo, Muming and Liu, Longqi and Naumann*, Robert K. and Xu*, Chun and Wang*, Zhenlong and Xu*, Xun and Liu*, Shiping}, title = {Genomic evolution reshapes cell-type diversification in the amniote brain}, journal = {Developmental Cell}, year = {2025}, month = may, day = {13}, pages = {1-60}, volume = {60}, number = {}, doi = {10.1016/j.devcel.2025.04.014}, url = {https://www.sciencedirect.com/science/article/pii/S1534580725002527}, eprint = {https://www.sciencedirect.com/science/article/pii/S1534580725002527}, } -
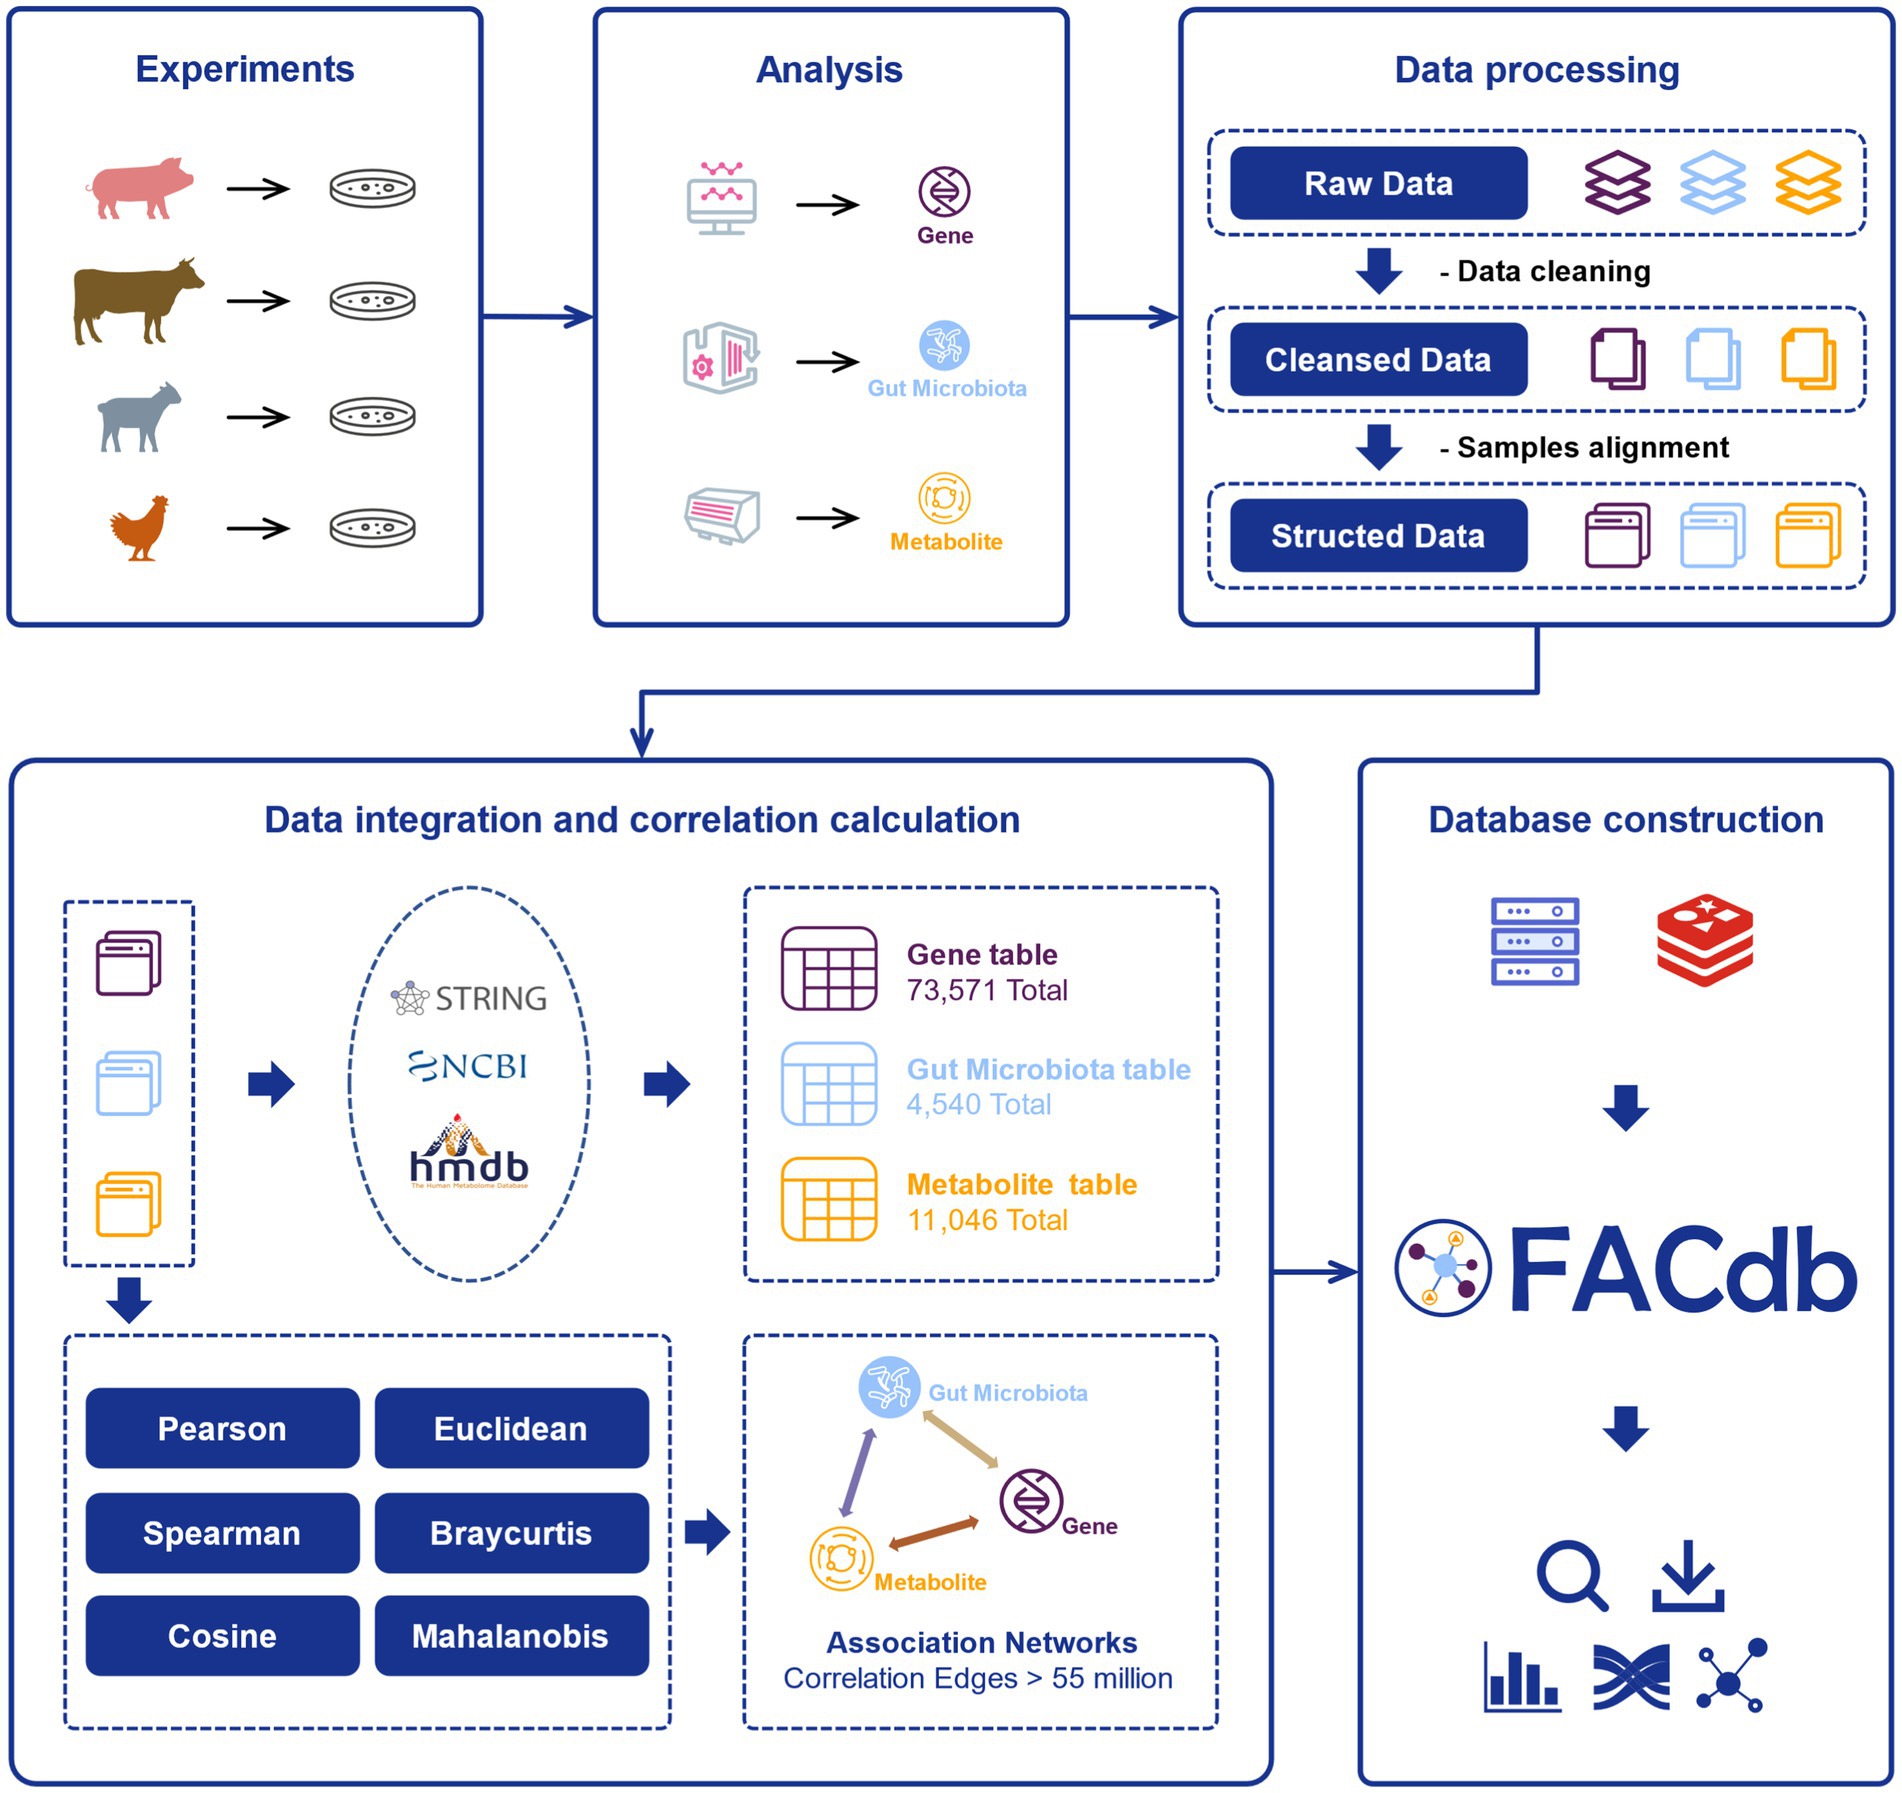 FACdb: a comprehensive resource for genes, gut microbiota, and metabolites in farm animalsZe Zhang† , Yang Li† , Di Zhang† , Shuai Chen* , Sien Lu , Kang Wang , Miao Zhou , Zehe Song , Qingcui Li* , Jie Yin* , and Xiaoping Liu*Frontiers in Microbiology, Mar 2025
Farm animals, including livestock and poultry, play essential economic, social, and cultural roles and are indispensable in human welfare. Farm Animal Connectome database (FACdb) is a comprehensive resource that includes the association networks among gene expression, gut microbiota, and metabolites in farm animals. Although some databases present the relationship between gut microbes, metabolites, and gene expression, these databases are limited to human and mouse species, with limited data for farm animals. In this database, we calculate the associations and summarize the connections among gene expression, gut microbiota, and metabolites in farm animals using six correlation or distance calculation (including Pearson, Spearman, Cosine, Euclidean, Bray–Curtis, and Mahalanobis). FACdb contains over 55 million potential interactions of 73,571 genes, 11,046 gut microbiota, and 4,540 metabolites. It provides an easy-to-use interface for browsing and searching the association information. Additionally, FACdb offers interactive visualization tools to effectively investigate the relationship among the genes, gut microbiota, and metabolites in farm animals. Overall, FACdb is a valuable resource for understanding interactions among gut microbiota, metabolites, and gene expression. It contributes to the further utilization of microbes in animal products and welfare promotion. Compared to mice, pigs or other farm animals share more similarities with humans in molecular, cellular, and organ-level responses, indicating that our database may offer new insights into the relationship among gut microbiota, metabolites, and gene expression in humans.
@article{10.3389/fmicb.2025.1557285,
  author = {Zhang†, Ze and Li†, Yang and Zhang†, Di and Chen*, Shuai and Lu, Sien and Wang, Kang and Zhou, Miao and Song, Zehe and Li*, Qingcui and Yin*, Jie and Liu*, Xiaoping},
  title = {FACdb: a comprehensive resource for genes, gut microbiota, and metabolites in farm animals},
  journal = {Frontiers in Microbiology},
  year = {2025},
  month = mar,
  day = {21},
  pages = {},
  volume = {16},
  number = {},
  doi = {10.3389/fmicb.2025.1557285},
  url = {https://www.frontiersin.org/journals/microbiology/articles/10.3389/fmicb.2025.1557285/full},
  eprint = {https://www.frontiersin.org/journals/microbiology/articles/10.3389/fmicb.2025.1557285/full},
}


2024
-
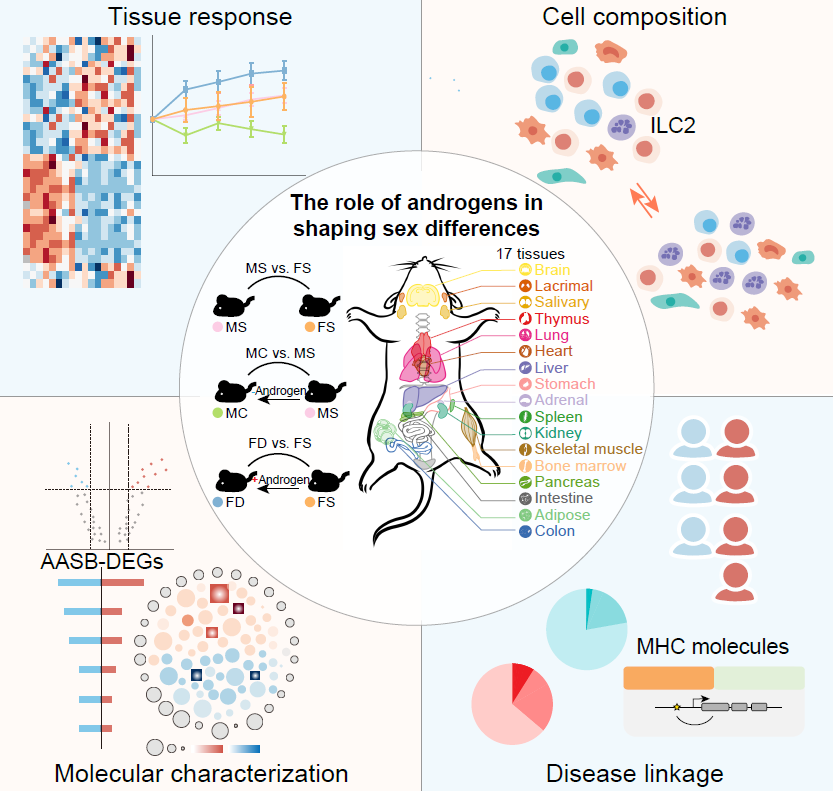 Sex differences orchestrated by androgens at single-cell resolutionFei Li† , Xudong Xing† , Qiqi Jin† , Xiang-Ming Wang† , Pengfei Dai† , Ming Han† , Huili Shi , Ze Zhang , Xianlong Shao , Yunyi Peng , Yiqin Zhu , Jiayi Xu , Dan Li , Yu Chen , Wei Wu , Qiao Wang , Chen Yu* , Luonan Chen* , Fan Bai* , and Dong Gao*Nature, Apr 2024
Sex differences in mammalian complex traits are prevalent and are intimately associated with androgens1-7. However, a molecular and cellular profile of sex differences and their modulation by androgens is still lacking. Here we constructed a high-dimensional single-cell transcriptomic atlas comprising over 2.3 million cells from 17 tissues in Mus musculus and explored the effects of sex and androgens on the molecular programs and cellular populations. In particular, we found that sex-biased immune gene expression and immune cell populations, such as group 2 innate lymphoid cells, were modulated by androgens. Integration with the UK Biobank dataset revealed potential cellular targets and risk gene enrichment in antigen presentation for sex-biased diseases. This study lays the groundwork for understanding the sex differences orchestrated by androgens and provides important evidence for targeting the androgen pathway as a broad therapeutic strategy for sex-biased diseases.
@article{doi:10.1038/s41586-024-07291-6,
  author = {Li†, Fei and Xing†, Xudong and Jin†, Qiqi and Wang†, Xiang-Ming and Dai†, Pengfei and Han†, Ming and Shi, Huili and Zhang, Ze and Shao, Xianlong and Peng, Yunyi and Zhu, Yiqin and Xu, Jiayi and Li, Dan and Chen, Yu and Wu, Wei and Wang, Qiao and Yu*, Chen and Chen*, Luonan and Bai*, Fan and Gao*, Dong},
  title = {Sex differences orchestrated by androgens at single-cell resolution},
  journal = {Nature},
  volume = {},
  number = {},
  pages = {1-8},
  year = {2024},
  month = apr,
  day = {10},
  doi = {10.1038/s41586-024-07291-6},
  url = {https://www.nature.com/articles/s41586-024-07291-6},
  eprint = {https://www.nature.com/articles/s41586-024-07291-6},
}

2023
-
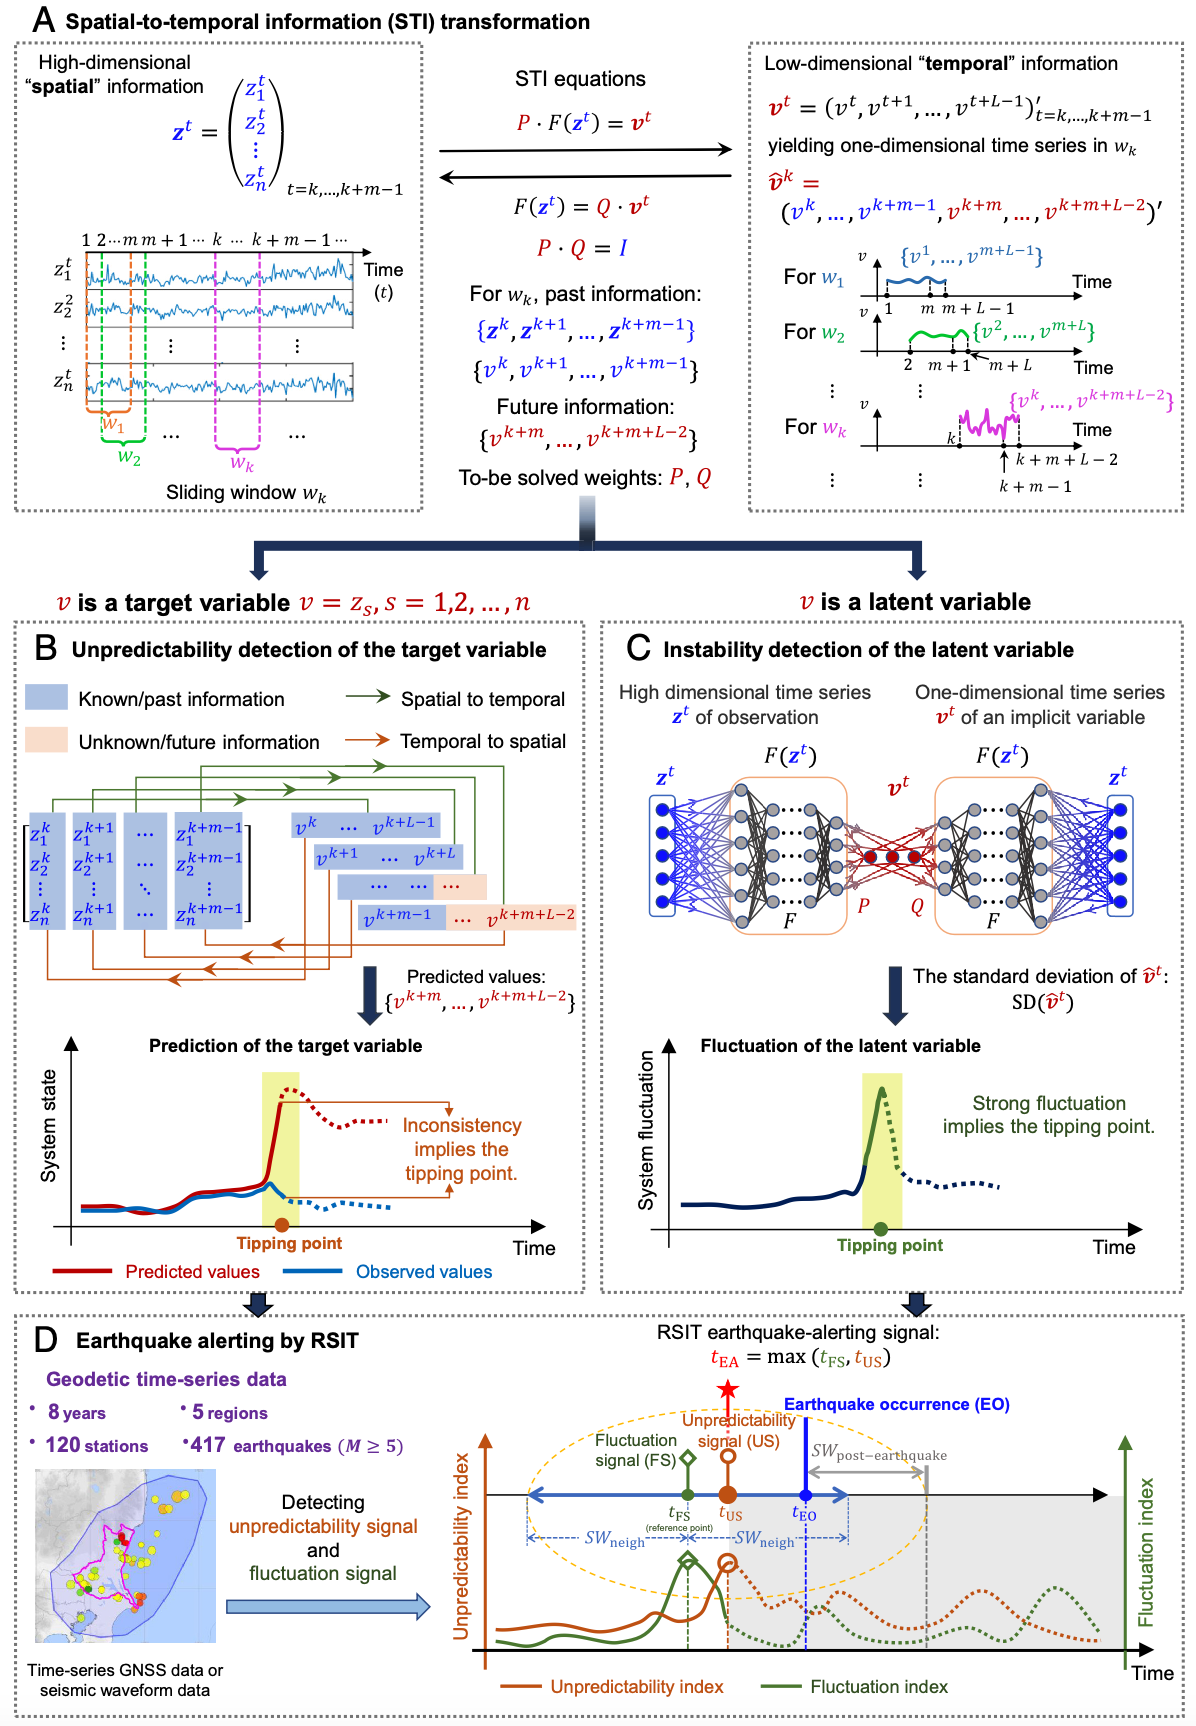 Earthquake alerting based on spatial geodetic data by spatiotemporal information transformation learningYuyan Tong† , Renhao Hong† , Ze Zhang , Kazuyuki Aihara , Pei Chen* , Rui Liu* , and Luonan Chen*Proceedings of the National Academy of Sciences (PNAS), Sep 2023
Alerting for imminent earthquakes is particularly challenging due to the high nonlinearity and nonstationarity of geodynamical phenomena. In this study, based on spatiotemporal information (STI) transformation for high-dimensional real-time data, we developed a model-free framework, i.e., real-time spatiotemporal information transformation learning (RSIT), for extending the nonlinear and nonstationary time series. Specifically, by transforming high-dimensional information of the global navigation satellite system into one-dimensional dynamics via the STI strategy, RSIT efficiently utilizes two criteria of the transformed one-dimensional dynamics, i.e., unpredictability and instability. Such two criteria contemporaneously signal a potential critical transition of the geodynamical system, thereby providing early-warning signals of possible upcoming earthquakes. RSIT explores both the spatial and temporal dynamics of real-world data on the basis of a solid theoretical background in nonlinear dynamics and delay-embedding theory. The effectiveness of RSIT was demonstrated on geodynamical data of recent earthquakes from a number of regions across at least 4 y and through further comparison with existing methods.
@article{doi:10.1073/pnas.2302275120, author = {Tong†, Yuyan and Hong†, Renhao and Zhang, Ze and Aihara, Kazuyuki and Chen*, Pei and Liu*, Rui and Chen*, Luonan}, title = {Earthquake alerting based on spatial geodetic data by spatiotemporal information transformation learning}, journal = {Proceedings of the National Academy of Sciences (PNAS)}, volume = {120}, number = {37}, pages = {e2302275120}, year = {2023}, month = sep, day = {05}, doi = {10.1073/pnas.2302275120}, url = {https://www.pnas.org/doi/abs/10.1073/pnas.2302275120}, eprint = {https://www.pnas.org/doi/pdf/10.1073/pnas.2302275120}, } -
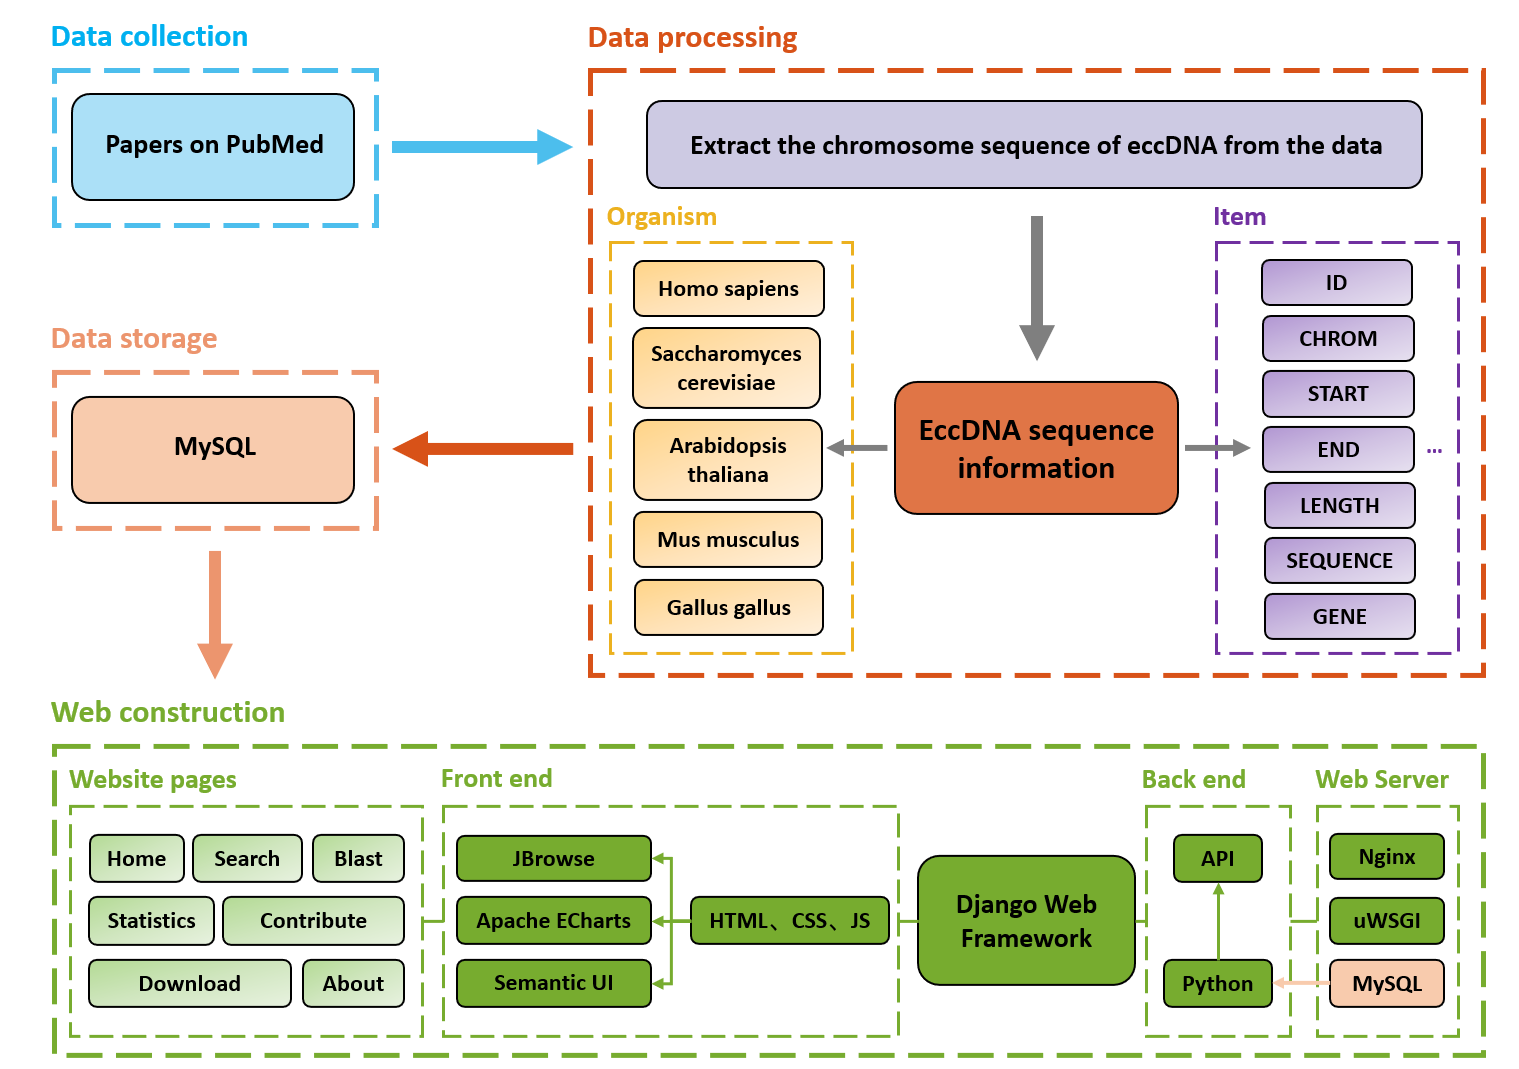 TeCD: The eccDNA Collection Database for extrachromosomal circular DNAJing Guo† , Ze Zhang† , Qingcui Li , Xiao Chang* , and Xiaoping Liu*BMC Genomics, Jan 2023
Background Extrachromosomal circular DNA (eccDNA) is a kind of DNA that widely exists in eukaryotic cells. Studies in recent years have shown that eccDNA is often enriched during tumors and aging, and participates in the development of cell physiological activities in a special way, so people have paid more and more attention to the eccDNA, and it has also become a critical new topic in modern biological research. Description We built a database to collect eccDNA, including animals, plants and fungi, and provide researchers with an eccDNA retrieval platform. The collected eccDNAs were processed in a uniform format and classified according to the species to which it belongs and the chromosome of the source. Each eccDNA record contained sequence length, start and end sites on the corresponding chromosome, order of the bases, genomic elements such as genes and transposons, and other information in the respective sequencing experiment. All the data were stored into the TeCD (The eccDNA Collection Database) and the BLAST (Basic Local Alignment Search Tool) sequence alignment function was also added into the database for analyzing the potential eccDNA sequences. Conclusion We built TeCD, a platform for users to search and obtain eccDNA data, and analyzed the possible potential functions of eccDNA. These findings may provide a basis and direction for researchers to further explore the biological significance of eccDNA in the future.
@article{doi:10.1186/s12864-023-09135-5,
  author = {Guo†, Jing and Zhang†, Ze and Li, Qingcui and Chang*, Xiao and Liu*, Xiaoping},
  title = {TeCD: The eccDNA Collection Database for extrachromosomal circular DNA},
  journal = {BMC Genomics},
  volume = {24},
  number = {1},
  pages = {47},
  year = {2023},
  month = jan,
  day = {27},
  doi = {10.1186/s12864-023-09135-5},
  url = {https://bmcgenomics.biomedcentral.com/articles/10.1186/s12864-023-09135-5},
  eprint = {https://bmcgenomics.biomedcentral.com/articles/10.1186/s12864-023-09135-5},
}

